Flumazenil is a benchmark benzodiazepine antagonist used to understand what happens at GABA-A receptors when antagonists bind, especially at the benzodiazepine site. This article explains, in receptor and kinetic terms, how flumazenil and similar compounds interact with GABA-A receptors, what “antagonism” means at the level of binding and gating, and how these interactions are studied in lab and preclinical systems.
Content overview
Introduction
GABA-A receptors and benzodiazepine binding sites
What “antagonist binding” means in receptor kinetics
Flumazenil as a model benzodiazepine antagonist
Association and dissociation: on-rates and off-rates for flumazenil
Competitive interactions between flumazenil and benzodiazepine agonists
Allosteric coupling: how flumazenil affects GABA gating
Kinetic models used to study flumazenil and GABA-A receptors
Longer-term effects: trafficking, plasticity, and receptor turnover
Experimental methods for quantifying benzodiazepine antagonist kinetics
Summary
FAQ
References
Introduction
Benzodiazepine antagonists are ligands that bind to the benzodiazepine site on GABA-A receptors without producing the same positive allosteric modulation as classical benzodiazepine agonists. Flumazenil is the best-characterized example and is widely used as a reference compound in receptor-binding, electrophysiological, and imaging studies.
Understanding what happens when flumazenil binds requires a kinetic perspective: how quickly it associates and dissociates from the receptor, how it competes with agonists and inverse agonists at the benzodiazepine site, and how its occupancy changes the probability that GABA will open the chloride channel. These processes are typically studied in vitro or in animal models, using controlled concentrations and time courses that allow receptor-level phenomena to be isolated from broader physiological variables.
GABA-A receptors and benzodiazepine binding sites
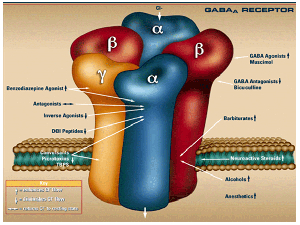
GABA-A receptors are pentameric ligand-gated chloride channels, usually assembled from combinations of α, β, and γ subunits. The orthosteric binding sites for GABA lie at the interfaces between α and β subunits, while benzodiazepine ligands—including flumazenil—bind at an allosteric site formed by the interface between an α subunit (α1, α2, α3, or α5) and the γ2 subunit.
When GABA binds and activates the receptor, the channel opens and chloride ions flow through, producing fast inhibitory postsynaptic currents. Benzodiazepine-site ligands modulate this process; classical agonists increase the receptor’s apparent affinity for GABA or enhance channel open probability, while inverse agonists decrease it. Antagonists such as flumazenil occupy the same allosteric pocket but, in many experimental systems, show little intrinsic effect on gating in the absence of other benzodiazepine-site ligands.
What “antagonist binding” means in receptor kinetics
In receptor kinetics, “antagonist binding” usually refers to a ligand that binds to a receptor without activating it, preventing other ligands from binding and exerting their effects. At the benzodiazepine site, flumazenil behaves primarily as a competitive antagonist: it binds reversibly and competes with agonists and inverse agonists for occupancy of the same site.
Kinetically, this can be described by association and dissociation rate constants. The association rate constant (kon_\text{on}on) and dissociation rate constant , which reflects affinity. A benzodiazepine antagonist with a relatively smallwill remain bound longer and occupy a high fraction of available receptors at a given concentration, effectively reducing access for other ligands that share the site.
Flumazenil as a model benzodiazepine antagonist
Flumazenil is an imidazobenzodiazepine derivative designed to retain high affinity for the benzodiazepine site while lacking strong positive modulatory activity. In many preparations containing α1β2γ2 or related receptors, flumazenil competes effectively with benzodiazepine agonists and inverse agonists and shows only weak intrinsic effects on channel function when applied alone.
Because flumazenil can be formulated at high purity and characterized by multiple analytical methods, it has become a standard reagent for defining benzodiazepine-site affinity and kinetics. In binding assays, flumazenil is often used as a “cold” competitor against radiolabeled benzodiazepine analogs, and in electrophysiology it is applied to reverse benzodiazepine-induced potentiation of GABA-evoked currents.
Association and dissociation: on-rates and off-rates for flumazenil
When flumazenil is applied to a preparation of GABA-A receptors, ligand–receptor complexes form at a rate governed by the product of kon_\text{on}on and the free flumazenil concentration. As complexes form, the reverse process—dissociation at rate koff_\text{off}off—removes ligand from the receptor, and the system approaches equilibrium.
In saturation binding experiments, this dynamic is reflected in how quickly a maximal occupancy level is reached and how readily binding is reversed by washout or by adding a competing ligand. Flumazenil’s relatively fast association and dissociation make it suitable for kinetic analyses where time resolution matters, such as stopped-flow spectroscopy or rapid perfusion electrophysiology. These experiments often show that flumazenil can rapidly occupy and vacate the benzodiazepine site, allowing researchers to track receptor responses to transient changes in occupancy.
Competitive interactions between flumazenil and benzodiazepine agonists
From a kinetic perspective, a major effect of benzodiazepine antagonists is to shift the apparent concentration–response curves of agonists and inverse agonists. In the presence of flumazenil, a benzodiazepine agonist requires higher concentrations to achieve the same level of receptor occupancy because both ligands share the benzodiazepine site. This manifests as a rightward shift in agonist dose–response curves in binding and functional experiments. Different benzodiazepine antagonists, such as flumazenil and experimental compounds /comparing-benzodiazepine-antagonists-in-the-laboratory-flumazenil-vs-experimental-compounds/, show variations in binding affinity and kinetics.
In radioligand binding assays, increasing flumazenil concentrations compete with a fixed amount of radiolabeled benzodiazepine analog, reducing specific binding in a concentration-dependent manner. The shape of this inhibition curve, together with known tracer parameters, allows estimation of flumazenil’s Ki_ii (inhibition constant). Functionally, in electrophysiology, flumazenil can reverse benzodiazepine-induced increases in GABA-evoked currents; the extent and speed of reversal depend on both ligands’ kinetic parameters and their relative concentrations.
Allosteric coupling: how flumazenil affects GABA gating
The benzodiazepine site does not directly open the channel; instead, it modulates the relationship between GABA binding and channel gating. Agonists stabilize receptor conformations with higher GABA efficacy, while inverse agonists stabilize conformations with lower efficacy. Antagonists like flumazenil tend to occupy the site without strongly favoring either conformation, though subtle effects can occur depending on subunit composition and experimental conditions.
In single-channel and macroscopic current recordings, this means that flumazenil alone often produces minimal change in opening frequency or mean open time relative to GABA alone. However, when GABA and a benzodiazepine agonist are present, adding flumazenil shifts the conformational equilibrium back toward the GABA-alone state by displacing the agonist from its site. Kinetic models incorporate this by allowing benzodiazepine-site occupancy to alter transition rates between closed, open, and desensitized states; flumazenil binding constrains those transitions to the GABA-only regime.
Kinetic models used to study flumazenil and GABA-A receptors
Several classes of kinetic models are used to interpret how benzodiazepine antagonists behave at GABA-A receptors:
Simple competitive binding models, which treat the benzodiazepine site as a single binding pocket with multiple ligands competing for occupancy.
Allosteric models, where binding at the benzodiazepine site modifies GABA binding affinity or gating transitions, such as Monod–Wyman–Changeux (MWC)-type schemes adapted for ligand-gated ion channels.
Single-channel Markov models, which define discrete open, closed, and desensitized states with transition rates that depend on both GABA and benzodiazepine-site occupancy.
Using these frameworks, experimental time courses of binding and current responses under various combinations of GABA, benzodiazepine agonists, and flumazenil can be fit to derive rate constants, conformational equilibria, and allosteric coupling factors. This helps distinguish pure competitive antagonism from more complex behavior such as weak inverse modulation or subtype-dependent effects.
Longer-term effects: trafficking, plasticity, and receptor turnover
While receptor kinetics often focus on millisecond-to-second timescales, prolonged exposure to benzodiazepine antagonists can influence slower processes such as receptor trafficking and turnover. Some in vitro studies have shown that extended flumazenil exposure alters surface expression of certain GABA-A receptor configurations, affecting internalization and degradation pathways.
For example, experiments in recombinant cell systems have reported changes in surface levels of δ-containing GABA-A receptors after hours of flumazenil treatment, interpreted as altered endocytosis or lysosomal targeting. At more classical αβγ receptors, chronic antagonist exposure has been associated in some models with changes in benzodiazepine binding site density or expression of GABA-binding subunits. These findings suggest that even ligands with minimal acute efficacy can, over time, modulate receptor population dynamics via transcriptional, translational, or trafficking mechanisms.
Experimental methods for quantifying benzodiazepine antagonist kinetics
A variety of techniques are used to investigate what happens when flumazenil and related antagonists bind:
Radioligand binding assays, including saturation and competition experiments, and can be adapted for kinetic (time-resolved) measurements.
Patch-clamp electrophysiology, in whole-cell or single-channel formats, measures changes in GABA-evoked currents when benzodiazepine agonists and flumazenil are applied alone or together. Rapid solution exchange allows time courses of antagonist onset and washout to be quantified.
Stopped-flow and rapid-mixing fluorescence assays, which follow conformational or binding-dependent signals on millisecond timescales, can capture association and dissociation rates for benzodiazepine-site ligands.
Positron emission tomography (PET) with radiolabeled flumazenil analogs provides in vivo estimates of binding potential and apparent kinetic parameters at the tissue level, using compartmental modeling to infer association and dissociation behavior under tracer conditions.
Imaging and biochemical assays of receptor trafficking, including surface biotinylation and confocal microscopy, probe longer-term changes in surface expression and internalization after chronic exposure to antagonists like flumazenil.
Summary
When benzodiazepine antagonists such as flumazenil bind to GABA-A receptors, they occupy the benzodiazepine site without strongly shifting the receptor toward higher or lower GABA efficacy in many experimental systems. Kinetically, they compete with agonists and inverse agonists for the same site, altering the probabilities that these modulators can bind and change channel behavior. The key parameters are association and dissociation rates, equilibrium affinities, and allosteric coupling factors that link benzodiazepine-site occupancy to GABA binding and gating.
Flumazenil’s well-characterized binding profile, rapid yet reversible kinetics, and relatively neutral functional effect make it a powerful reference ligand for studying benzodiazepine-site receptor kinetics. Its use spans radioligand binding, electrophysiology, PET imaging, and receptor trafficking studies, all framed in preclinical or receptor-focused contexts. Over longer timescales, chronic exposure to benzodiazepine antagonists can influence receptor expression and plasticity, illustrating that kinetic binding events are embedded in broader regulatory networks governing GABA-A receptor populations.
FAQ
What does a benzodiazepine antagonist like flumazenil do at the receptor level?
It binds reversibly to the benzodiazepine site on GABA-A receptors, competing with agonists and inverse agonists for occupancy. In many experimental systems, it has little intrinsic effect on GABA-evoked currents when applied alone, but it prevents other benzodiazepine-site ligands from enhancing or reducing those currents.
How is affinity of flumazenil for the benzodiazepine site measured?
Affinity is typically measured in radioligand binding assays, where increasing concentrations of flumazenil compete with a fixed concentration of a radiolabeled benzodiazepine analog. The resulting inhibition curves are analyzed to obtain inhibition constants and, with appropriate models, dissociation constants.
Can flumazenil have any direct effect on GABA-A receptor currents?
Some recombinant receptor studies have reported weak intrinsic modulation by flumazenil, depending on subunit composition and conditions. However, compared with classical agonists or inverse agonists, these effects are generally small, and flumazenil is widely used as a functional antagonist in preclinical work.
How do receptor kinetics change when flumazenil is added to a benzodiazepine agonist?
Flumazenil reduces the fraction of receptors occupied by the agonist by competing for the benzodiazepine site. This leads to reduced potentiation of GABA-evoked currents and rightward shifts in dose–response curves for the agonist, without necessarily changing GABA affinity itself.
Why are association and dissociation rates important for benzodiazepine antagonists?
On- and off-rates determine how quickly antagonists can block or unblock the benzodiazepine site when concentrations change. Fast kinetics allow rapid reversal of agonist effects in experimental systems, while slow dissociation would prolong antagonism even after extracellular levels decline.
Can benzodiazepine antagonists affect receptor expression over time?
Yes. Although their acute effects are dominated by competitive binding, prolonged exposure in laboratory models can alter receptor trafficking, internalization, or degradation, leading to changes in surface receptor levels and benzodiazepine binding site density. These longer-term effects are typically studied over hours to days in vitro or in animal models.
References
Flumazenil modulation of GABA-A receptor trafficking and surface expression in recombinant systems
https://pmc.ncbi.nlm.nih.gov/articles/PMC4696891/
PET studies using [11C]flumazenil to examine changes in benzodiazepine-site binding associated with altered GABA levels
https://journals.plos.org/plosone/article?id=10.1371%2Fjournal.pone.0032443
Experimental work characterizing intrinsic flumazenil activity and its effects on GABA-A receptor currents
https://pmc.ncbi.nlm.nih.gov/articles/PMC5326685/
Reviews and chapters describing flumazenil as a competitive benzodiazepine antagonist at α[1–3,5]βγ GABA-A receptors and summarizing kinetic behavior
https://www.sciencedirect.com/topics/pharmacology-toxicology-and-pharmaceutical-science/flumazenil
Studies addressing transcriptional and translational mechanisms in GABA-A receptor regulation under chronic flumazenil exposure
https://www.sciencedirect.com/science/article/abs/pii/S0168010208000795
Structural and dynamic analyses of GABA-A receptor modulators, including benzodiazepine-site ligands, in cryo-EM and computational models
https://pmc.ncbi.nlm.nih.gov/articles/PMC9357065/